Making mouse predictions#
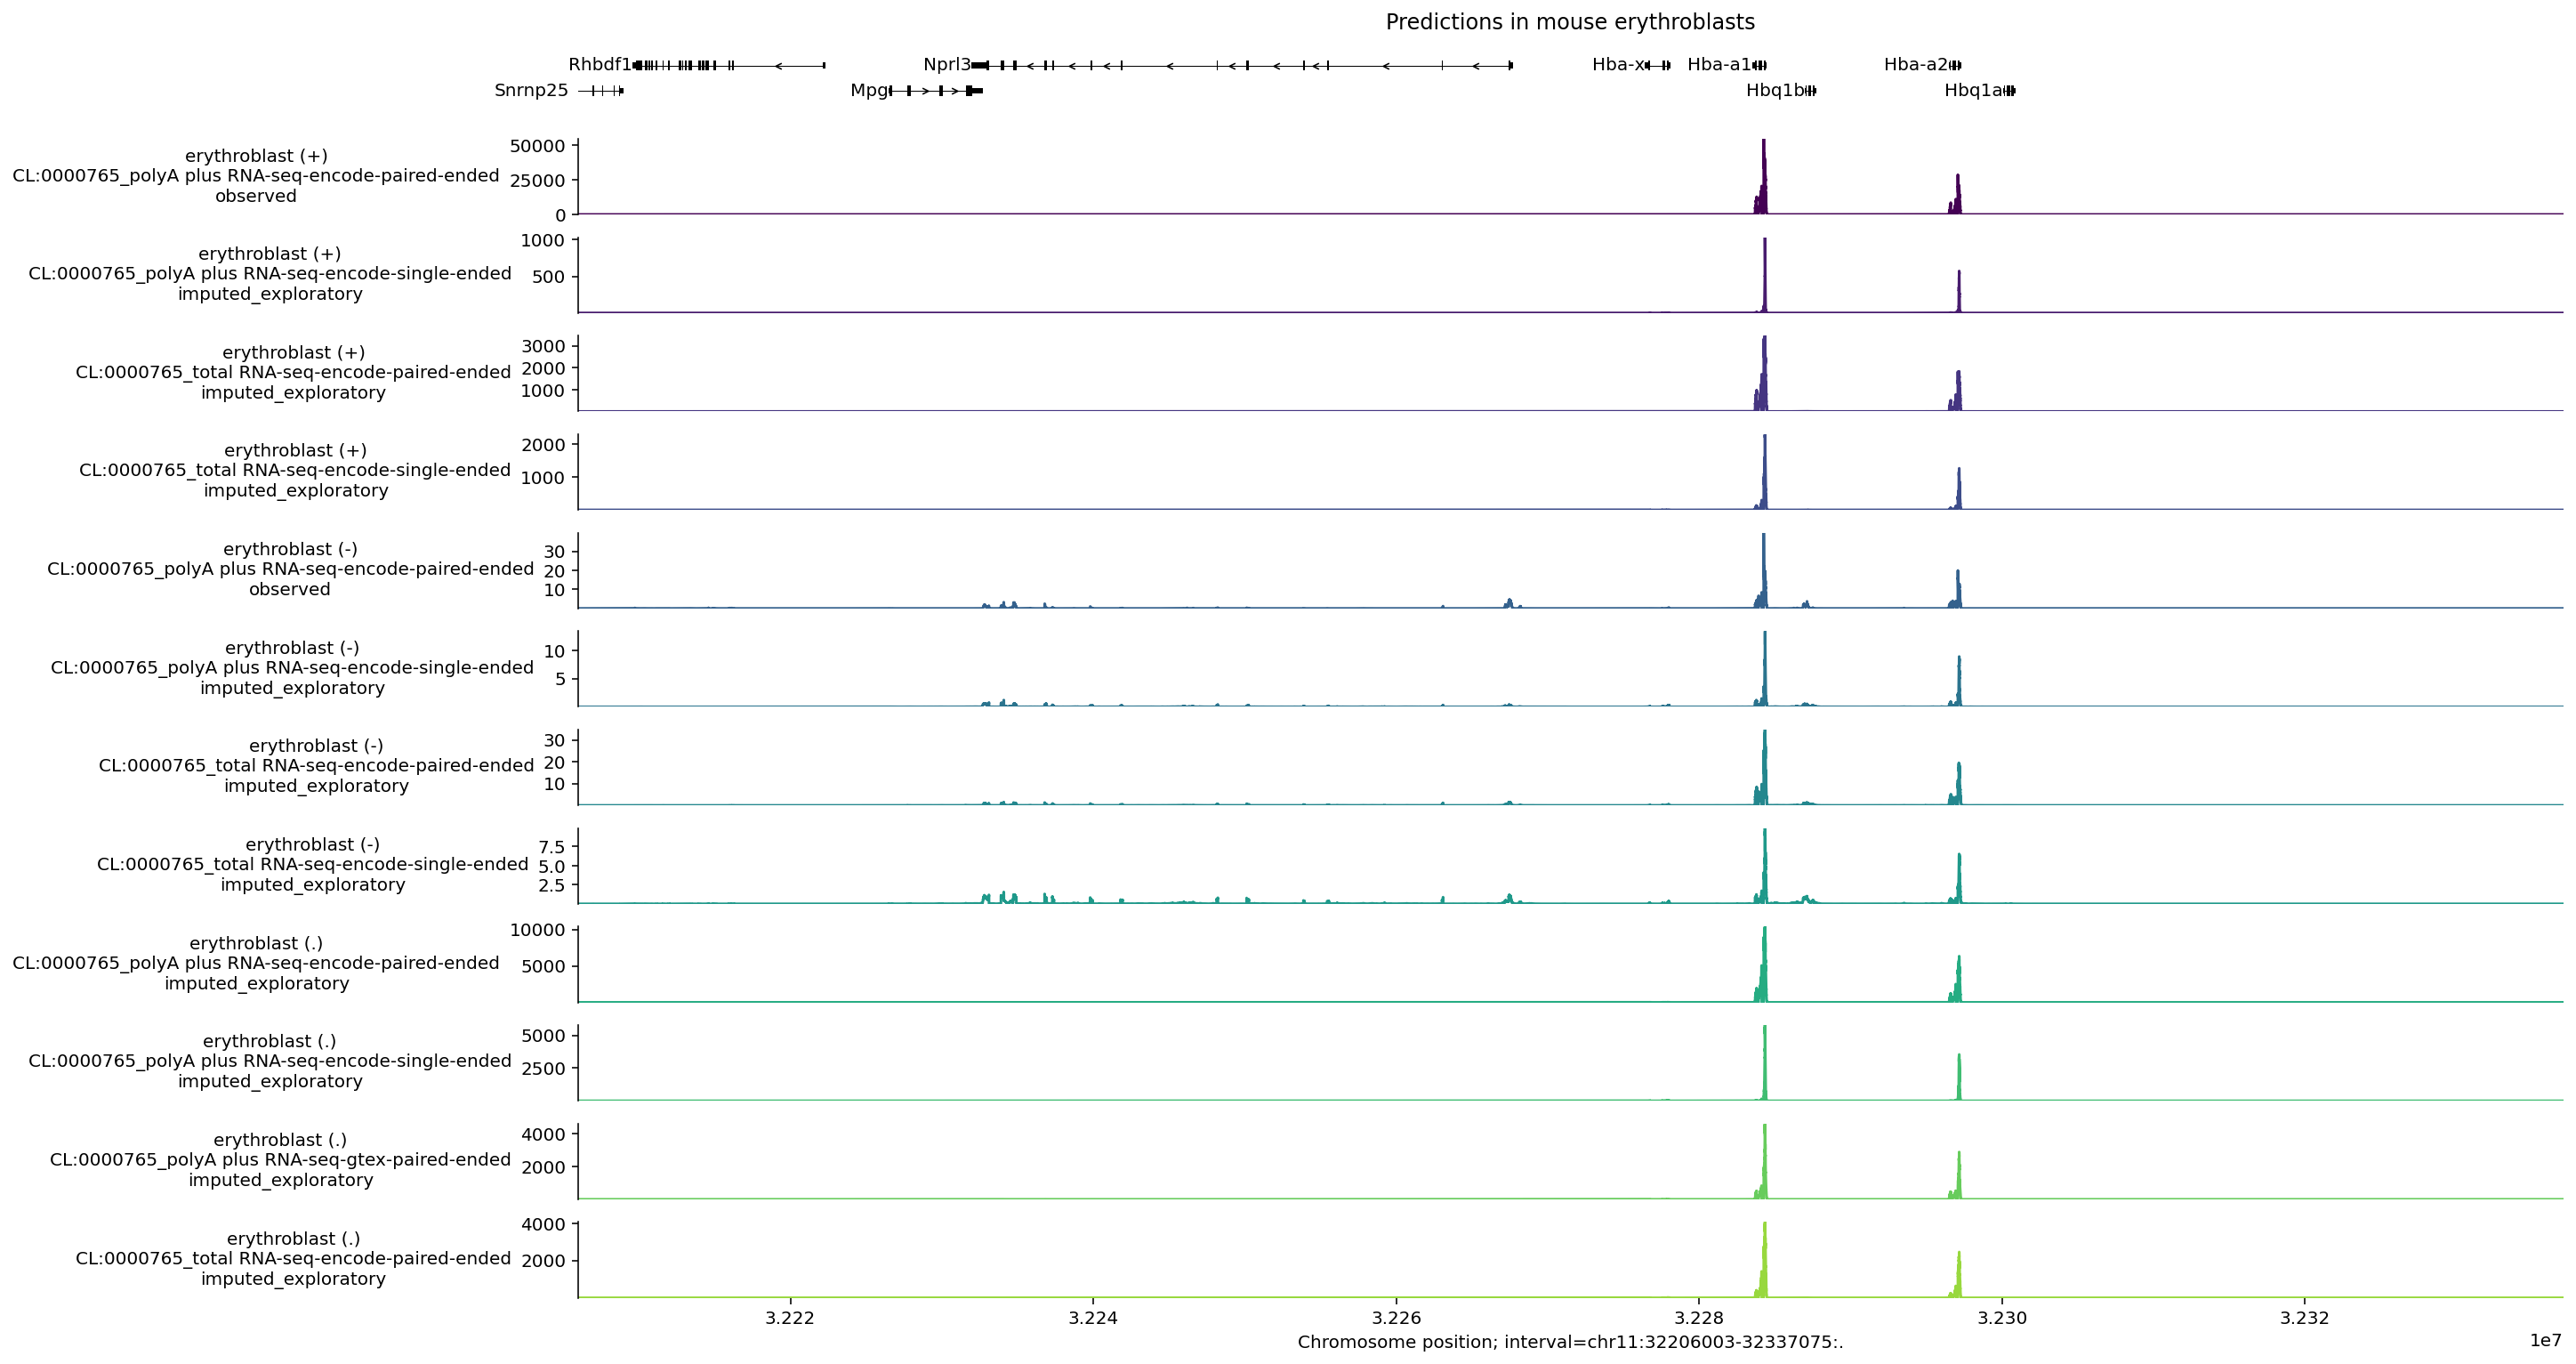
AlphaGenome 2 makes predictions for mouse (Mus musculus) as well as human. This tutorial predicts RNA-seq coverage across the mouse alpha-globin cluster in erythroblasts.
Key activities:
Load GENCODE vM23 (mm10) gene annotations
List mouse reference conditions with
supported_conditionsCall
predict_intervalwithorganism=dna_client.Organism.MUS_MUSCULUS
Set up#
# @title Install AlphaGenome

# @markdown Run this cell to install AlphaGenome.
from IPython.display import clear_output
! pip install alphagenome
clear_output()
# @title Imports
from alphagenome import colab_utils
from alphagenome.data import gene_annotation
from alphagenome.data import genome
from alphagenome.data import transcript as transcript_utils
from alphagenome.models.v2 import dna_client
from alphagenome.visualization import plot_components
import matplotlib.pyplot as plt
import pandas as pd
Store your API key in Colab Secrets for persistent access across sessions (see Installation), then initialize the client:
dna_model = dna_client.create(colab_utils.get_api_key())
Load GENCODE vM23 (mm10) gene annotations and build a TranscriptExtractor for the longest protein-coding transcript per gene (see GENCODE GTF format):
# The GTF file contains information on the location of all transcripts.
# Note that we use genome assembly mm10 for mice.
gtf = pd.read_feather(
    'https://storage.googleapis.com/alphagenome/reference/gencode/'
    'mm10/gencode.vM23.annotation.gtf.gz.feather'
)

# Get the longest transcript per gene
gtf_transcripts = gene_annotation.filter_protein_coding(gtf)
gtf_transcripts_longest = gene_annotation.filter_to_longest_transcript(
    gtf_transcripts
)

# Set up transcript extractors using the information in the GTF file.
transcript_extractor = transcript_utils.TranscriptExtractor(
    gtf_transcripts_longest
)
Make mouse predictions#
Other tutorials use the default organism, human (Organism.HOMO_SAPIENS). For mouse predictions, pass organism=dna_client.Organism.MUS_MUSCULUS to both supported_conditions and predict_interval, and pass a mouse ontology CURIE via condition=. Expression conditioning (cell_conditioning=) is supported for human only:
dna_model.supported_conditions(organism=dna_client.Organism.MUS_MUSCULUS)
| condition | biosample_type | biosample_name | atac | cage | dnase | rna_seq | chip_histone | chip_tf | splice_sites | splice_site_usage | splice_junctions | contact_maps | sc_atac | sc_gene_expression | methylation | |
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| 0 | CL:0000038 | primary_cell | erythroid progenitor cell | imputed_supported | imputed_exploratory | imputed_supported | imputed_supported | imputed_supported | imputed_exploratory | imputed_exploratory | imputed_exploratory | imputed_exploratory | imputed_exploratory | observed | observed | imputed_supported |
| 1 | CL:0000048 | primary_cell | multi fate stem cell | imputed_supported | imputed_exploratory | imputed_supported | imputed_supported | imputed_supported | imputed_exploratory | imputed_exploratory | imputed_exploratory | imputed_exploratory | imputed_exploratory | observed | observed | imputed_supported |
| 2 | CL:0000136 | primary_cell | adipocyte | imputed_supported | imputed_exploratory | observed | imputed_supported | imputed_supported | imputed_exploratory | imputed_exploratory | imputed_exploratory | imputed_exploratory | imputed_exploratory | imputed_supported | imputed_supported | imputed_supported |
| 3 | CL:0000187 | primary_cell | muscle cell | imputed_supported | imputed_exploratory | imputed_supported | observed | observed | observed | observed | observed | observed | imputed_exploratory | imputed_supported | imputed_supported | imputed_exploratory |
| 4 | CL:0000556 | primary_cell | megakaryocyte | imputed_supported | imputed_exploratory | imputed_supported | observed | observed | imputed_exploratory | observed | observed | observed | imputed_exploratory | imputed_supported | imputed_supported | imputed_exploratory |
| ... | ... | ... | ... | ... | ... | ... | ... | ... | ... | ... | ... | ... | ... | ... | ... | ... |
| 715 | UBERON:0008345 | tissue | ileal epithelium | imputed_exploratory | observed | imputed_exploratory | imputed_exploratory | imputed_exploratory | imputed_exploratory | imputed_exploratory | imputed_exploratory | imputed_exploratory | imputed_exploratory | imputed_exploratory | imputed_exploratory | imputed_exploratory |
| 716 | UBERON:0008800 | tissue | parietal endoderm | imputed_supported | imputed_exploratory | imputed_supported | imputed_supported | imputed_supported | imputed_exploratory | imputed_exploratory | imputed_exploratory | imputed_exploratory | imputed_exploratory | observed | observed | imputed_supported |
| 717 | UBERON:0008945 | tissue | extraembryonic endoderm | imputed_supported | imputed_exploratory | imputed_supported | imputed_supported | imputed_supported | imputed_exploratory | imputed_exploratory | imputed_exploratory | imputed_exploratory | imputed_exploratory | observed | observed | imputed_supported |
| 718 | UBERON:0010230 | tissue | eyeball of camera-type eye | imputed_exploratory | observed | imputed_exploratory | imputed_exploratory | imputed_exploratory | imputed_exploratory | imputed_exploratory | imputed_exploratory | imputed_exploratory | imputed_exploratory | imputed_exploratory | imputed_exploratory | imputed_exploratory |
| 719 | UBERON:0012314 | tissue | embryonic facial prominence | observed | imputed_exploratory | observed | observed | observed | imputed_exploratory | imputed_exploratory | imputed_exploratory | imputed_exploratory | imputed_exploratory | imputed_supported | imputed_supported | imputed_supported |
720 rows × 16 columns
Call predict_interval for a mouse genomic interval around the alpha-globin cluster (chr11:32206003-32337075) in mouse erythroblasts (CL:0000765; use dna_client.SEQUENCE_LENGTH_1MB for highest accuracy):
interval = genome.Interval.from_str('chr11:32206003-32337075').resize(
    dna_client.SEQUENCE_LENGTH_100KB
)

output = dna_model.predict_interval(
    interval=interval,
    organism=dna_client.Organism.MUS_MUSCULUS,
    requested_outputs=[dna_client.OutputType.RNA_SEQ],
    condition='CL:0000765',
)

output.rna_seq.values.shape
(131072, 12)
# Extract transcripts
transcripts = transcript_extractor.extract(interval)

# Plot
plot_components.plot(
    components=[
        # Add transcripts to plot
        plot_components.TranscriptAnnotation(transcripts),
        # Add RNA output
        plot_components.Tracks(
            output.rna_seq,
            ylabel_template=(
                '{biosample_name} ({strand})\n{name}\n{imputation_mode}'
            ),
        ),
    ],
    interval=interval,
    title='Predictions in mouse erythroblasts',
)

plt.show()